Diagnostics
IVD & Diagnostic Validation
IVD organizations sit at the intersection of device manufacturing, reagent production and high-volume laboratory operations — which means the validation portfolio spans process, equipment, instrument and informatics work simultaneously.
Reagent and consumable manufacturing
- Formulation, filling and lyophilization process validation
- Cold chain, storage and stability equipment qualification
- Cleaning validation for shared reagent equipment
- Packaging, labeling and serialization systems
Instrument systems and automation
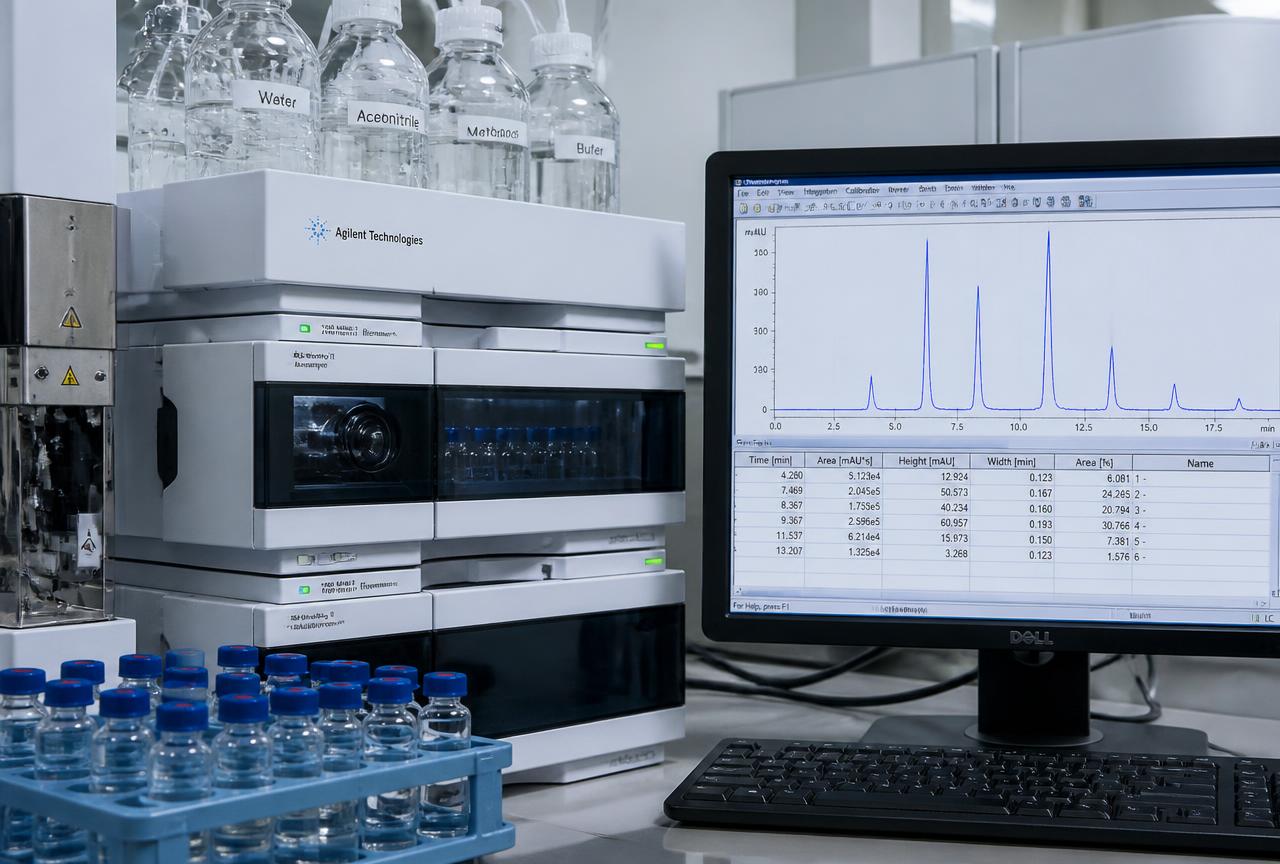
Diagnostic platforms combine mechanical handling, fluidics, detection and software. Qualification of the manufacturing and test equipment that builds and verifies these systems has to account for the measurement chain that ultimately determines whether a produced instrument meets specification.
Informatics and data integrity
- LIMS and middleware in high-throughput operations
- Result calculation, flagging and release logic
- Interfaces to instruments, EHR and reporting systems
- Access control, audit trails and electronic result review
- Complete data capture for repeats, QC failures and recalibrations
Separate IVD performance evidence from manufacturing assurance
EU Regulation (EU) 2017/746 defines IVD scope through intended purpose, including qualifying software used for examination of specimens derived from the human body. General laboratory and research-use-only products are excluded unless specifically intended for IVD examination. Determine the product's regulatory scope before selecting a validation framework.
IVD performance evaluation addresses scientific validity, analytical performance and clinical performance under Annex XIII. Manufacturing process validation, instrument qualification and laboratory-system assurance support controlled production and data handling, but do not substitute for device performance evidence. In the US, evaluate applicable device requirements and QMSR separately from the EU IVDR framework.
- Trace intended purpose and performance claims to appropriate evidence
- Challenge calculation, flags, QC failure handling and result transmission
- Assess device software separately from production/QMS software
- Control reagent, instrument, software and interface changes against intended use
Primary references
- FDA — Process Validation: General Principles and Practices
- FDA — Computer Software Assurance for Production and Quality Management System Software
- FDA — 21 CFR Part 11, Electronic Records; Electronic Signatures
- FDA — Data Integrity and Compliance With Drug CGMP
- FDA — General Principles of Software Validation
- EUR-Lex — Regulation (EU) 2017/746 on in vitro diagnostic medical devices
Information published on ValidationEngineering.com is educational and informational. It is not legal or regulatory advice and is not a guarantee of regulatory compliance or of any inspection outcome. Organizations remain responsible for their own quality decisions.
Next step
Validating diagnostic manufacturing or laboratory systems?
Describe your scope and timeline.